Dynamic changes to the epigenome play a critical role in establishing and maintaining cellular phenotype during differentiation, but little is known about the normal methylomic differences that occur between functionally distinct areas of the brain. In this project we are characterizing intra- and inter-individual methylomic variation across whole blood and multiple regions of the brain from multiple donors using MeDIP-seq and Illumina 450K arrays.
We observe distinct tissue-specific patterns of DNA methylation, with a highly significant over-representation of tissue-specific differentially methylated regions (TS-DMRs) observed at intragenic CpG islands and low CG density promoters. A large proportion of TS-DMRs are located near genes that are differentially expressed across brain regions and TS-DMRs are significantly enriched near genes involved in functional pathways related to neurodevelopment and neuronal differentiation, including BDNF, BMP4, CACNA1A, CACA1AF, EOMES, NGFR, NUMBL, PCDH9, SLIT1, SLITRK1 and SHANK3.
Although between-tissue variation in DNA methylation greatly exceeds between-individual differences within any one tissue, we observe that some inter-individual variation is reflected across brain and blood, indicating that peripheral tissues may have some utility in epidemiological studies of complex neurobiological phenotypes. This study reinforces the importance of DNA methylation in regulating cellular phenotype across tissues, and highlights genomic patterns of epigenetic variation across functionally distinct regions of the brain, providing a resource for the epigenetics and neuroscience research communities.
Data from this project can be viewed here or via the NIH Epigenomics Atlas.
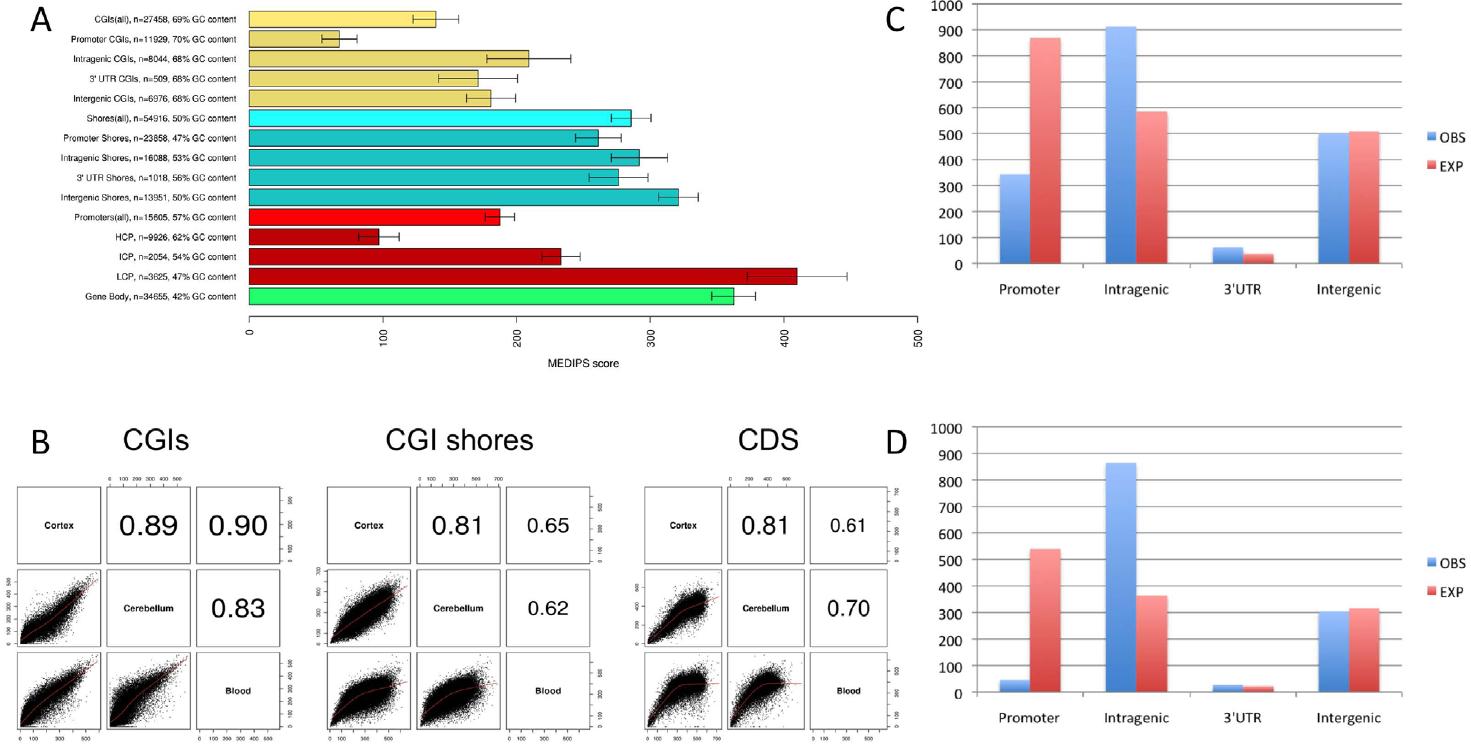
Although DNA methylation at CGIs is relatively conserved across tissues, intragenic CGIs are dramatically over-represented and promoter CGIs under-represented in the most tissue-variable CGIs. (a) Average DNA methylation values calculated by MEDIPS from MeDIP-seq data for all annotated gene features: CGIs (yellow), CGI shores (blue), gene promoters (red) and CDSs (green). DNA methylation is lower in promoter CGIs compared to intragenic, 3' UTR and intergenic CGIs. CGI shores are characterized by higher DNA methylation than CGIs, with less location-dependent variation. Promoter DNA methylation shows a strong inverse correlation with GC density, with LCPs showing a higher average level of DNA methylation than CDSs. Error bars represent standard error of the mean. (b) Although TS-DMRs are distributed across all feature types, there are marked differences in the between-tissue correlation of DNA methylation across each of the broad feature categories we examined, with CGIs being more correlated across cortex, cerebellum and blood than CGI shores or CDSs. (c, d) There is a highly significant enrichment of intragenic CGIs (P = 2 x 10-102) in analyses of CGI DMRs differentiating blood, cortex and cerebellum (c), and an even more dramatic enrichment (P = 1 x 10-246) in comparisons between cortex and cerebellum (d). EXP, expected; OBS, observed.